Plot a normalized hic map with annotations
Source:R/shaman_plot_map.R
shaman_plot_map_score_with_annotations.Rdshaman_plot_map_score_with_annotations
shaman_plot_map_score_with_annotations(
  genome,
  points_score,
  interval_range,
  point_size = 0.1,
  misha_tracks = list(),
  mt_colors = getOption("shaman.track_colors"),
  mt_ylims = NULL,
  annotations = list(),
  a_colors = getOption("shaman.annotation_colors"),
  add_genes = TRUE,
  add_ideogram = TRUE,
  add_axis = TRUE,
  gene_stacking = "squish",
  gene_size = 0.7,
  track_size = 0.8,
  annotation_size = 0.7,
  fig_fn = "",
  fig_width = 900,
  fig_height = 5/6 * 900
)
Arguments
- genome
Name of reference genome (e.g. "hg19", "mm10")
- points_score
A dataframe containing the points (start1, start2) and their normalized score.
- interval_range
1D interval (chrom, start, end) specifying the region to plot
- point_size
Cex size of the points in the plot.
- misha_tracks
List of 1D track expressions (virtual tracks also supported) which can be extracted from.
- mt_colors
Array of colors, one for each misha_track, which will be used to plot each 1d track.
- mt_ylims
Y-axis limits for each misha track. If not provided, generated automatically based on the data to be displayed in the region.
- annotations
List of gintervals highlighting annotated regions.
- a_colors
Array of colors, one for each annotation set.
- add_genes
Boolean flag indicating whether to show gene annotations.
- add_ideogram
Boolean flag indicating whether to add a chromosome ideogram.
- add_axis
Boolean flag indicating whether to show the chromosomal axis.
- gene_stacking
Describes the viewing option of the genes track. Can be either "squish", or "dense".
- gene_size
Size of gene annotation view.
- track_size
Size of track view.
- annotation_size
Size of annotation view.
- fig_fn
Name of png file to output to. Empty string will cause the figure to be plotted to the current device.
- fig_width
Width in pixels of output png figure.
- fig_height
Height in pixels of output png figure.
Value
No return value, called for the plot. Needs the Bioconductor package Gviz.
Details
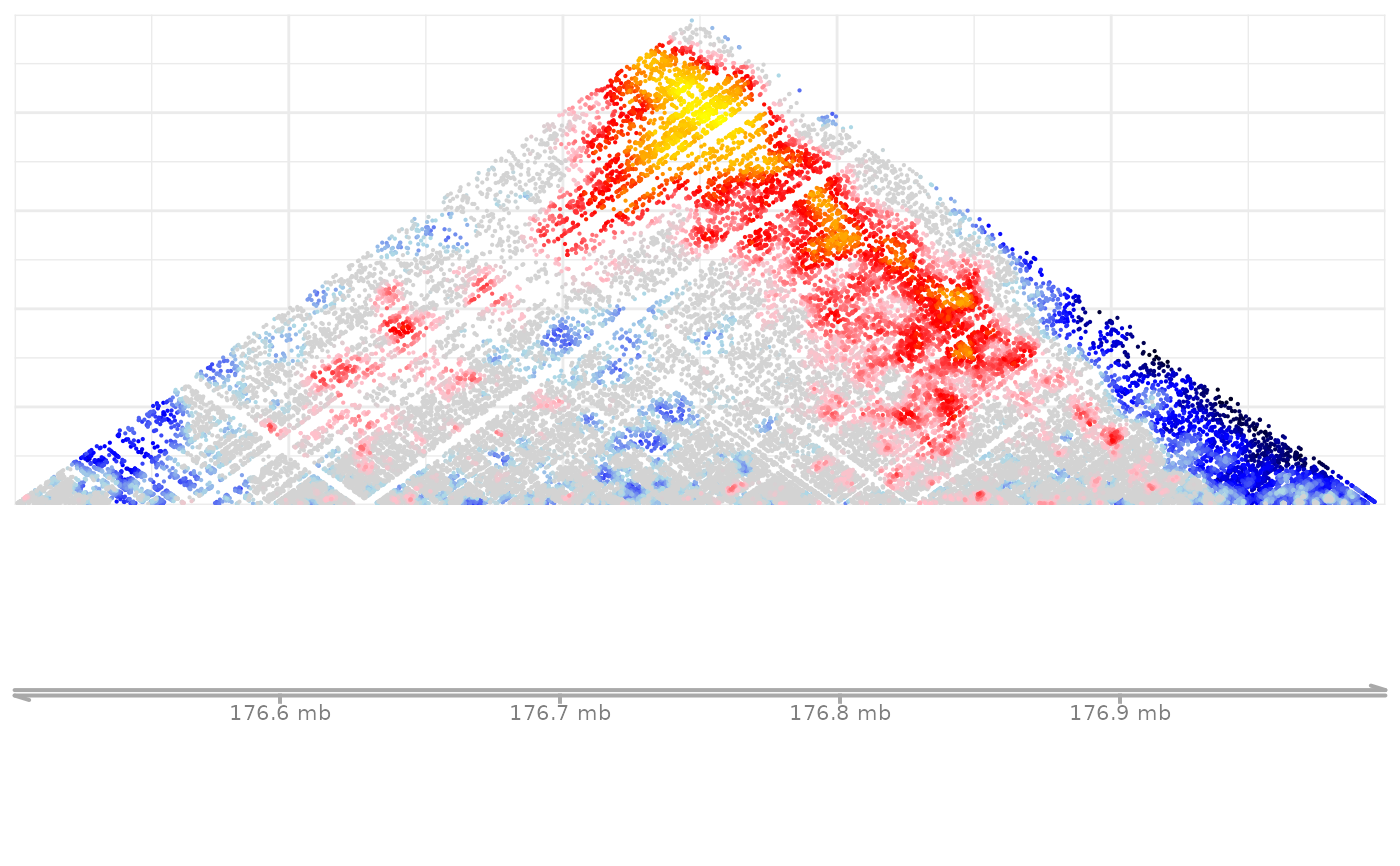
Plots observed hic contact matrix color-coded by normalized scores, aligned with linear data such as gene annotations, rna, chromatin, transcription factor binding, etc.).
Examples
library(misha)
gsetroot(shaman_get_test_track_db())
points <- gextract("hic_score", gintervals.2d(2, 176.5e06, 177e06, 2, 176.5e06, 177e06),
colnames = "score"
)
# \donttest{
# loading Gviz takes a few seconds
if (requireNamespace("Gviz", quietly = TRUE)) {
shaman_plot_map_score_with_annotations("hg19", points, gintervals(2, 176.5e06, 177e06),
add_genes = FALSE, add_ideogram = FALSE
)
}
 # }
# gene annotations and the ideogram are downloaded from UCSC
if (interactive() && requireNamespace("Gviz", quietly = TRUE)) {
shaman_plot_map_score_with_annotations("hg19", points, gintervals(2, 176.5e06, 177e06))
}